• Psychedelic Drugs Welcome Guest
    View threads about
    Posting RulesBluelight Rules
    PD's Best Threads Index
    Social ThreadSupport Bluelight
    Psychedelic Beginner's FAQ
  • PD Moderators: Esperighanto | JackARoe |

Phenethylamines How can I purify 2C-B at home?

Honestly this is a great question which I truly don't know the answer. My guess* is that the 2C-H that was used for bromination had other impurities and those impurities got brominated too. They then get trapped in the crystal matrix of the 2C-B itself, but as they probably don't form a salt with HBr or HCL, they tend to be very soluble in organic solvents while 2C-B salts are not very* soluble in isopropyl. This is why I would say a wash is the easiest way one could improve the quality of 2C-B at home without much gear or risk. Would agree the antisolvent (water/acetone) method is trickier.

If it was residual bromine itself, I would think airing it out would remove the color as bromine tends to evaporate super easily. I've never smelled bromine in 2C-B and the tan 2C-B that I have seen, I've seen it get even darker over time. That makes me think brominated side products are present and are even more prone to oxidation than the 2C-B itself. Once the 2C-B is white, it never goes dark again IME.

I also want to say here that while 2C-B that is cut likely does exist, I would say most of the dirty 2C-B we all see is not washed enough or not recrystallized and the impurities are generally synthesis by products. @xdrc is probably right about an anhydrous isopropyl alcohol recrystallization possibly being superior to the water/acetone antisolvent crystallization. I would be careful though as it involves heating a flamnable solvent. Once all the 2C-B is entirely dissolved, simply let it return to room temperature slowly, the slower the better. Filtering when it is hot could not hurt either.

If it is CUT 2C-B, other strategies may be more effective.

Ok so I have some more questions.

Can you chemistry-understanding people take a look at the common route(s) of making 2C-B and determine what these bromine side compounds might be?

OR

Could it be another compound from earlier in the synth that makes its way up to the 2C-H step?

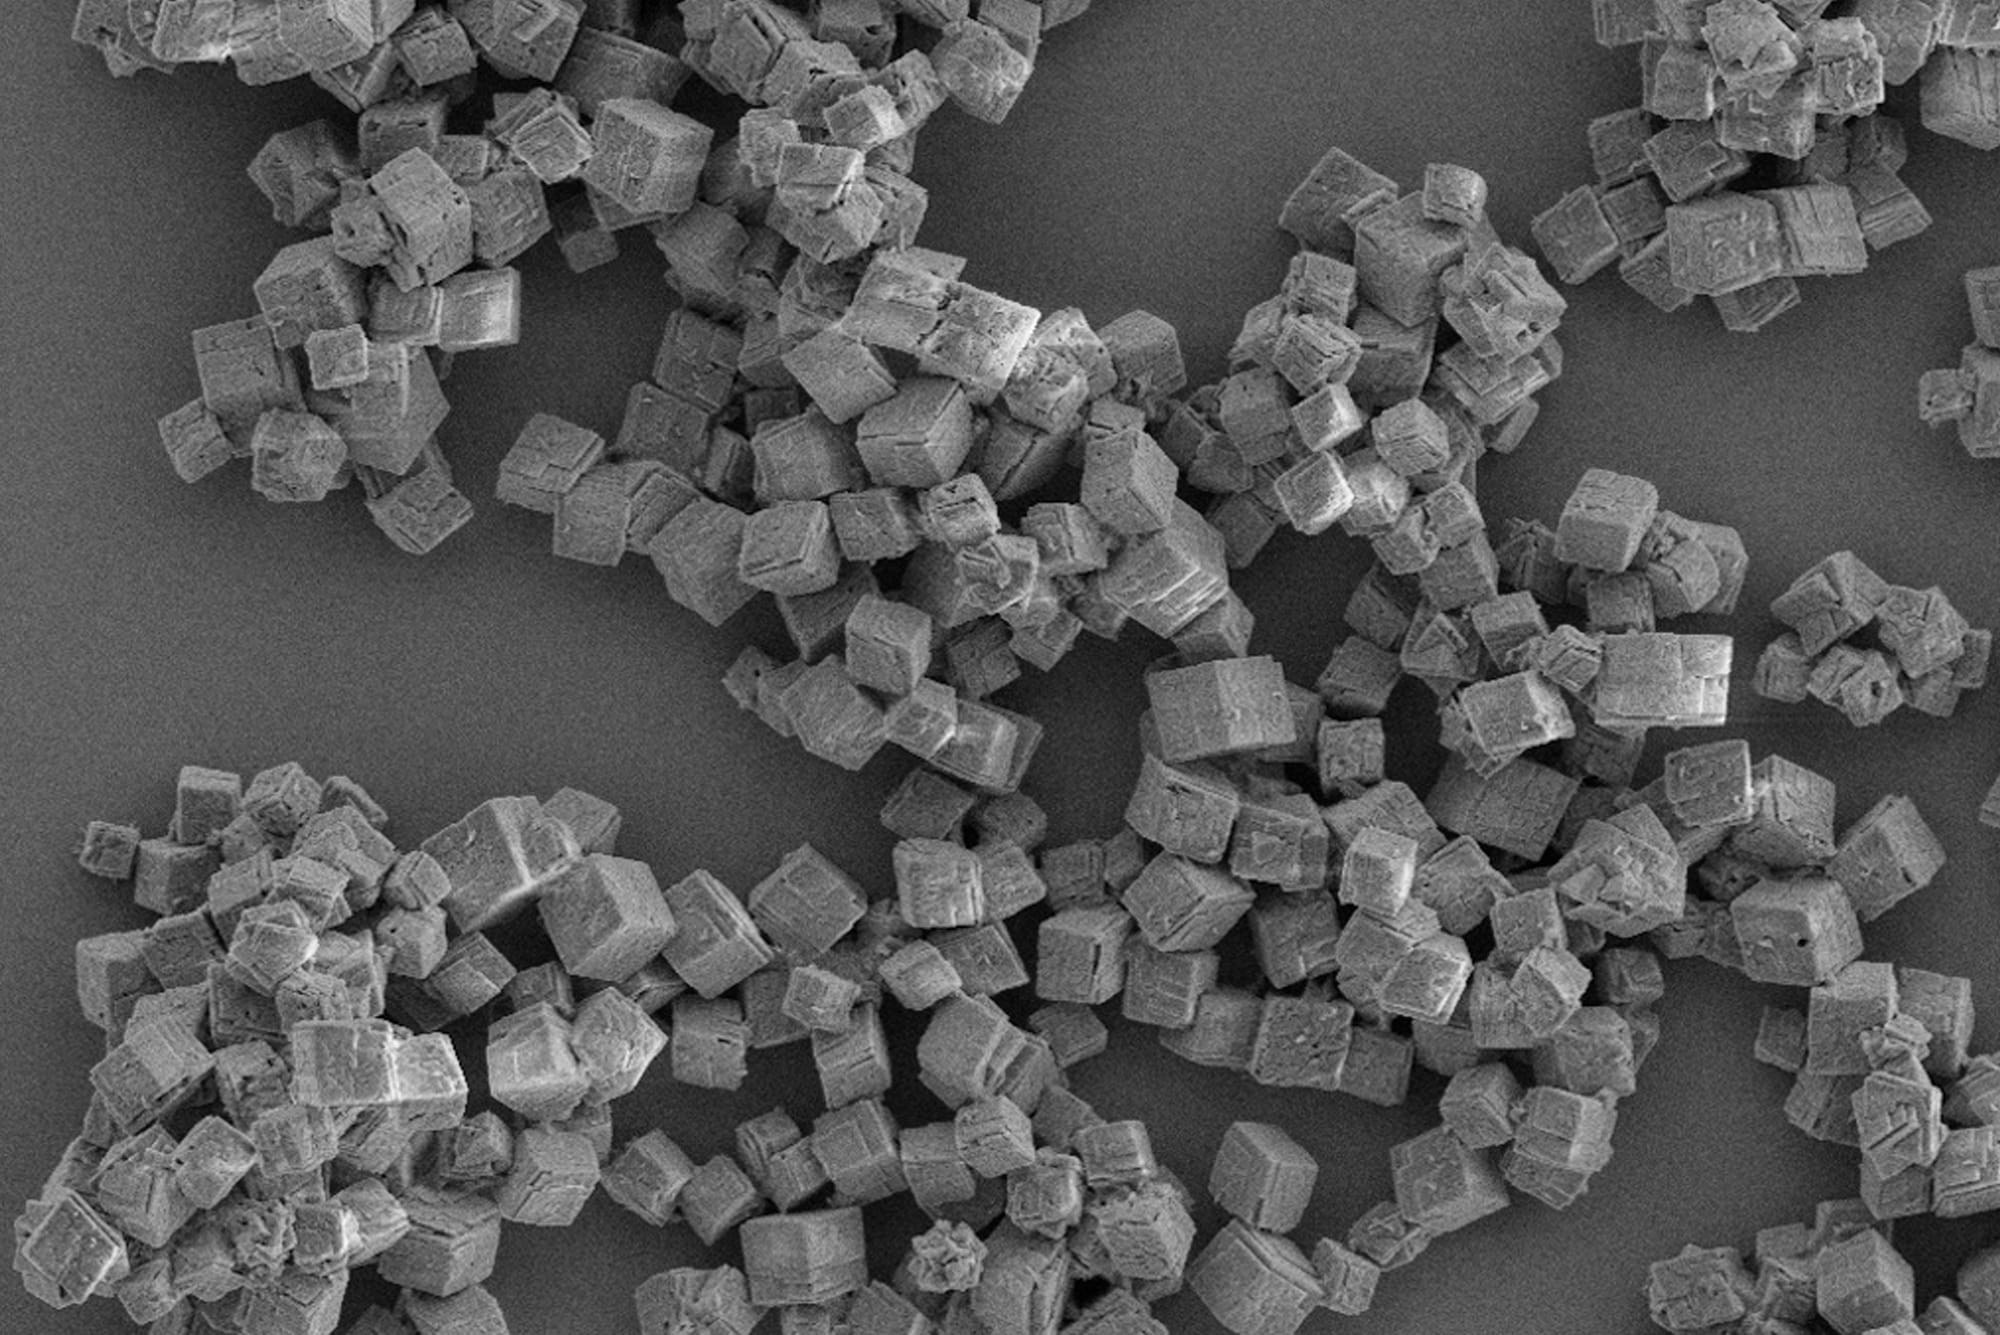
Regarding crystallization, where I am in Europe most of the 2C-B is brown powder or a pill. I assume that the chemist synths are rapid precipitation and that they're not doing anything slow at all. It all seems to come out of the Netherlands? so I assume it's being manufactured by the MDMA cartels who disdainfully do a side run of 2C-B when they're ordered to once a year. The MDMA crystal rocks are out of control perfect so obviously they know how to crystallize. They must really not give a shit about the 2c-b.

Has anyone successfully homegrown cool crystals of 2c-b? Any notes?
ima take a shot at this, from the rip there is no way to know at the quantum lattice formation level what byproducts are in there without lab equipment
However, looking at the side chains substituents, and functional groups: you could get among other things, Excess, unreacted elemental bromine or acidic bromination byproducts causing orangish tanning
During reduction stages: nitrostyrene byproducts
Oxidized freebase impurities
2-4, 2-6 brominated Aromatic locations instead of 2-5
Brominating agents can cause minor cleavage of the sensitive methoxy -OCH3 group
Residual benzaldehyde

MDMA is water loving molecule, dissolves easily in water, warm alcohols, or acetone-alcohol mixes and flexible methylenedioxy rings allow nice lattice stacking
However, 2C-B, is hydrophobic and the moment 2C-B freebase oil meets hydrochloric acid, the resulting salt desperately wants to crash out violently and instantly as a precipitated powder. The molecules lock together so fast that they trap passing impurities and solvents inside their structure (from research papers stuff)


Highly Soluble Molecule (MDMA) + Heat ---> Dissolves Completely
---> Cools SLOWLY over days
---> Molecules line up perfectly
---> Large Macro-Crystals (Rocks)

Poorly Soluble Molecule (2C-B) + Acid ---> Instantly Hates Solvent
---> Crashes out violently in seconds
---> Traps impurities & solvent
---> Microcrystalline Tan Powder

water gets trapped in those crystals for mdma lot easier than they ever would in 2CB


and furthermore, reading: "2C-B is active in tiny doses (meaning a single small batch goes a very long way), clandestine labs want to use the fastest method possible: dissolve the 2C-H freebase in a solvent like ether or acetone, bubble HCl gas through it, and instantly filter out the brown/tan powder that floats to the top. They have no economic reason to slow down a process that is designed to be finished in minutes."

again ive not synthesized nor messed with this (never will), and this is all from papers and what bit i know of orgchem (the science all makes sense w/o even playing in a lab)
without synthesizing yet trying to explain this phenomenon:
1. Dissolving the HCl Salt in Water
Action: You take your phenylethylamine HCl salt (let's denote it as R-NH3+Cl-) and dissolve it in water.
Purpose: This is often done to get it into a medium where you can perform an acid-base extraction. The HCl salt is highly water-soluble, so it readily dissolves.

2. Basification of the Aqueous Solution
Action: You add a strong base, such as sodium hydroxide (NaOHNaOH) or potassium carbonate (K2CO3K2CO3), to the aqueous solution containing the R-NH3+Cl−. Ensuring the solution is sufficiently basic (e.g., pH 10-12).

The free base is typically less water soluble.
Depending on the specific phenylethylamine, it might:
Separate out as an immiscible oily layer on top or bottom of the aqueous solution.
Precipitate as a solid if its melting point is high enough and it's sufficiently insoluble in water.
Remain partially dissolved in the aqueous phase, but its concentration will be low compared to its solubility in organic solvents.

3. Extraction into an Organic Solvent ("Transfer to another organic solvent")
Action: You add an immiscible organic solvent to the basified aqueous solution containing the free base. Common choices include:
Diethyl ether: Low boiling point, good for heat-sensitive compounds.
Dichloromethane (DCM) / Chloroform: Denser than water, good for extracting into the bottom layer.
Toluene: Good for less polar free bases.
Mechanism: The free base (R-NH2) is typically much more soluble in the organic solvent than in water. When the two immiscible layers are mixed (in a separatory funnel), the free base preferentially migrates from the aqueous layer into the organic layer. Water-soluble impurities (like NaCl, excess NaOH) remain in the aqueous layer
Repetition: This extraction is usually performed multiple times (e.g., 2-3 times) with fresh portions of organic solvent to ensure maximum recovery of the free base.
Separation: The organic layer containing the dissolved free base is then separated from the aqueous layer.

4. Drying the Organic Solution
Action: The separated organic solution will inevitably contain some dissolved water. To prepare for crystallization of the HCl salt, this water must be removed. A drying agent (anhydrous MgSO4, Na2SO4, CaSO4, CaCl2) is added to the organic solution, stirred, and then filtered off.
Purpose: Water would react with the anhydrous HCl
HCl you're about to add, consuming it and potentially hindering salt formation or crystal growth.

5. Formation and Crystallization of the HCl Salt
Action: The dry organic solution of the free base (R-NH2) is treated with anhydrous HCl. This can be done by:
Bubbling dry HCl gas directly into the solution.
Adding a solution of HCl in a suitable anhydrous organic solvent (HCl in dry ether, HCl in isopropanol, HCl in acetone). The choice of solvent for the HCl solution should be one where the final HCl salt of the phenylethylamine is poorly soluble.

Crystallization: Since the phenylethylamine HCl salt is designed to be poorly soluble in the anhydrous organic solvent(s) used at this stage, it will precipitate out of solution as a solid.
To get well-formed crystals, the process should be slow and controlled (slow addition of HCl, gentle cooling if initially warmed, seeding if possible). Rapid addition or high concentrations can lead to amorphous precipitation (fine powder or oiling out).

Same for MDMA HCl. Look at meth off the birch reduction PSE base (another once freely avail compound one skip away from high intense energy, similar to phenylethylamine's one stop shop decarbox) that liquid ammonia once the HCl vapor from a lil sulfuric acid + salt hits that free base solution over there the compound by its nature can crystallize nicely, nevermind the -34C temp in that (lil off topic)


its a phenomenon not an exact perfected science - we know we can crash out precip stuff, sometimes we can cool crystallize stuff, sometimes it does it by its nature
 
Last edited:
from PiHKAL ("2,5-dimethoxyphenethylamine (2C-H) which picks up carbon dioxide from the air very quickly." - so the chances of having that particular form in ambient air laying around on a table or nightstand are slim to none, unless contained in appropriate storage solution/container
fyi that comment is sasha discussing the fact that 2C-H when it is a freebase and only when it is a freebase can react with CO2 in the air to create a carbonate salt. It's not ideal at all because it's very sticky/oily/waxy difficult to handle and probably won't participate in future halogenation reactions. You'd definitely be surprised by the way. Even mescaline FB can react with CO2 to create carbonate salts. I assume at least some other phenethylamines will too.
I may have a look at crystal formation in more detail, I can picture ostwaldt ripening working but, but not sure if by inclusions that changes the compound or not for a %, and if ostwaldt then mitigates the inclusions, guess it depends on the yield improvement v pain of ostwaldt.
@jmc31337 is right when he saying ostwaldt ripening not a direct inclusion removal mechanism (why do I think this is all copy pasted ai prompt) rather it is a way to grow huge pretty crystals. The bigger the crystal is, the more difficult it becomes to dismantle it to remove the inclusion. I never said it was a good way to purify anything I just said it was worth checking out if you interested in nerdy concepts and pretty crystals
 
Last edited:
thanks, I understand that a lot better now

I may have a look at crystal formation in more detail, I can picture ostwaldt ripening working but, but not sure if by inclusions that changes the compound or not for a %, and if ostwaldt then mitigates the inclusions, guess it depends on the yield improvement v pain of ostwaldt.

off down another learning rabbit hole...
Here is a non ai website with lots of good info and diagrams.


I actually had syrris come to my lab years ago and demo one of their crystallization units. Turns out they are really good at explaining without over complicating it.
 
Here is a non ai website with lots of good info and diagrams.


I actually had syrris come to my lab years ago and demo one of their crystallization units. Turns out they are really good at explaining without over complicating it.
thanks for this! very kind

I'll go have a read, looks very interesting
 
In the illicit field, time is against you, the market favors production of precipitated cocaine hcl over coke crystals - ya master that re-crystallization process im all ears btw (especially for how long it would take)
makes sense, the coke re-crystalisation I've come across is simply a "premium grade" wash marketed at the top end of the market, dissolve part of a brick on warm Ethanol and re-crystalise, then wash it with ethanol.

I have to say having just tried this for the 1st time recently I'm quite impressed by the quality of the coke it produces
Regarding crystallization, where I am in Europe most of the 2C-B is brown powder or a pill.
UK here so expect our 2C-B is sourced from similar places, the 2C-B HCL I last bought was off white grey. Any pills I've had tend to be shit tbh and underdosed, but finding HCL was a right pain in the UK, people seem to think pills are cool, the demand prefers them....
 
Here is a non ai website with lots of good info and diagrams.


I actually had syrris come to my lab years ago and demo one of their crystallization units. Turns out they are really good at explaining without over complicating it.
nucleation
solubility
all this was mentioned

The big thing i was leaning towards as far as science, is TEMP. Crystallization doesnt necessarily require cooled temps, or supersaturated solutions
The non ai article mentioned ice cream, and adipic acid
If ya wanna play with the facet of crystallization you can do tabe salt, sugar crystals. But drug solutions compared to ice cream salt and sugar are obviously different - and crystallization is still a phenomena (hence its not liq, solid, gas, crystal, forms of matter - crystals/diamonds are that form of matter phenomena)
 
fyi that comment is sasha discussing the fact that 2C-H when it is a freebase and only when it is a freebase can react with CO2 in the air to create a carbonate salt. It's not ideal at all because it's very sticky/oily/waxy difficult to handle and probably won't participate in future halogenation reactions. You'd definitely be surprised by the way. Even mescaline FB can react with CO2 to create carbonate salts. I assume at least some other phenethylamines will too.

@jmc31337 is right when he saying ostwaldt ripening not a direct inclusion removal mechanism (why do I think this is all copy pasted ai prompt) rather it is a way to grow huge pretty crystals. The bigger the crystal is, the more difficult it becomes to dismantle it to remove the inclusion. I never said it was a good way to purify anything I just said it was worth checking out if you interested in nerdy concepts and pretty crystals
wait, i can accept being wrong in certain areas and right in others - but i dont like being wrong when it comes to drugs

So i quoted the scientific article in dicta - i knew it was one of the shulgin team members didnt think it was needed since the quote leads ya right to their synthesis
What was i wrong about?
 
its honestly not a lot of effort in practice. I was just trying to be absolutely clear.

Crush up b, pour in glass, pour in acetone, swirl, pour through coffee filter.

Could one could avoid the filter losses by pipetting the acetone layer off and then just letting the rest evaporate? You don't get all the acetone but no filter losses. Could do a few times.

For the IPA RX how much alcohol should you use? Just enough to dissolve or enough to have a layer of IPA thar would submerge any crystals?
First ?: yeah for simple wash that would suffice, however, from the research the stuff is SLIGHTLY SOLUBLE/PARTIALLY SOLUBLE meaning you still dissolve a lil in acetone you could still pipette and let evap like ya said - that part is sound wash based upon compound solubility
Discarding the acetone wash stuff (which now holds the impurities dissolved within and some partial 2C) - hence i mentioned CUTS (why would ya need wash good off the lab line product if its done correct?) with phenylethylamine backbones (say aspartame, i dont see aspartame dissolving in acetone (also partially solub.) not trying re-spark the street-cut upsale arg again tho

And to the 2nd ?: Ill have to defer to someone else


Edit: free base forms however, a different story. My quick lookup shows FREE BASE FORMS of pehnylalenine, tyrasine, aspartame, sucrose are insoluable in acetone
Caffiene free base form is 2g/100mL acetone at room temp as far as solubility, however its salt form is not soluble
(re:street cuts)
 
Last edited:
fyi that comment is sasha discussing the fact that 2C-H when it is a freebase and only when it is a freebase can react with CO2 in the air to create a carbonate salt. It's not ideal at all because it's very sticky/oily/waxy difficult to handle and probably won't participate in future halogenation reactions. You'd definitely be surprised by the way. Even mescaline FB can react with CO2 to create carbonate salts. I assume at least some other phenethylamines will too.

@jmc31337 is right when he saying ostwaldt ripening not a direct inclusion removal mechanism (why do I think this is all copy pasted ai prompt) rather it is a way to grow huge pretty crystals. The bigger the crystal is, the more difficult it becomes to dismantle it to remove the inclusion. I never said it was a good way to purify anything I just said it was worth checking out if you interested in nerdy concepts and pretty crystals
lattice energy calculations via Born–Haber Cycle and ethalpy of formations for the chem nerds?
 
  • WTF
Reactions: iom
Could one could avoid the filter losses by pipetting the acetone layer off and then just letting the rest evaporate? You don't get all the acetone but no filter losses. Could do a few times.

For the IPA RX how much alcohol should you use? Just enough to dissolve or enough to have a layer of IPA thar would submerge any crystals?
The filter losses are referring to material that stick to the filter. Are you going to get every single nanogram off the filter once you place the 2c on to the filter? Probably not, so no you will not get around the filter losses. Plus with this pipette method, ANY amount of acetone that you leave to evaporate is just going to redeposit the acetone soluble impurities back on to the 2C-B. If you want to actually purify the 2C-B, allow the acetone (contains impurities) pass through the filter and use fresh acetone to rinse any dirty acetone off.

The 2nd question, for which jmc has no answer for is applicable to ALL recrystallization across chemistry. For a recrystallization to work it needs to be partially soluble in a solvent at a given temperature. If it can't go into solution, then you can never recrystallize. To minimize losses one uses the minimum amount of solvent possible to dissolve the crystal. Using excess solvent makes it that much more difficult to precipitate with temperature alone but excess solvent is going to make it that much harder for the molecules in solution to find the crystal lattice at the correct place to build upon and you're going to lose molecules in the solution. This is one of the most basic concepts of crystallization. You would think with all the Ai information at JMC's disposal, he would understand the basic concepts of crystallization. That shouldn't even be difficult to look up. In my opinion, the true skill is knowing how to ask the right question. This is what google ai says when I ask it "in crystallization why does one use the least amount of solvent". I'm posting it because I may not always be the best at explaining that which I already know.

Maximize Product Recovery (Yield)
  • Preventing loss in the mother liquor: Every solid has a small, non-zero solubility in a solvent, even when cooled down. If you use too much solvent, a large portion of your dissolved compound will remain trapped in the liquid (mother liquor) after cooling rather than precipitating out as crystals, resulting in a low yield. [1, 2]
  • Reaching supersaturation: A minimal amount of solvent ensures that the solution becomes heavily supersaturated upon cooling, which provides the necessary thermodynamic driving force for clean crystal formation. [1, 2]

Maintain Purity
  • Keeping impurities dissolved: Impurities are usually present in smaller amounts and are more soluble than the target compound. Using excess solvent means more impurities can stay dissolved or risk co-crystallizing, which compromises the purity of your final product. [1, 2]

Efficiency and Practicality
  • Faster cooling and handling: Smaller volumes require less thermal energy to heat and less time to cool down.
  • Reduced waste: Using less solvent generates less chemical waste, making the process more cost-effective and environmentally friendly. [1, 2, 3, 4]
----------------------------------------------------------------------
By the way I want to remind everyone the title of this thread.
Purification of 2C-B at home. Some key words are "at home"

To me that means the process is so basic that anyone can do with easily obtainable supplies. No lattice energy calculations required.
 
Last edited:
The filter losses are referring to material that stick to the filter. Are you going to get every single bit off the filter once you place the 2c on to the filter? Probably not, so no you will not get around the filter losses. Plus with this pipette method, ANY amount of acetone that you leave to evaporate is just going to redeposit any acetone soluble impurities back on to the 2C-B. If you want to actually purify the 2C-B, allow the acetone (contains impurities) pass through the filter and use fresh acetone to rinse any dirty acetone off.

The 2nd question for which jmc has no answer for is applicable to ALL recrystallization across chemistry. Think about it. For a recrystallization to work it needs to be partially soluble in a solvent at a given temperature. If it can't go into solution, then you can never recrystallize. Therefore the obvious answer is that to minimize losses one uses the minimum amount of solvent possible to dissolve the crystal. Using excess solvent makes it that much more difficult to precipitate with temperature alone but excess solvent is going to make it that much harder for the molecules in solution to find the crystal lattice at the correct place and you're going to lose molecules in the solution. This is one of the most basic concepts of crystallization.

You would think with all the Ai knowledge at JMC's disposal, he would understand the most basic concepts of crystallization.

By the way I want to remind everyone the title of this thread.

Purification. Of. 2C-B. At. Home.

Some key words are "At Home"

That means so basic that anyone can do with easily obtainable supplies. No lattice energy calculations required.
I did mention crystallography, even capitalized TEMP as what i felt was main factor towards crystallization
Then someone mentioned HCl and we discussed that
Then someone mentioned 2c-h and we ended up here
Lets assume two scenarios: someone hands you a liquid free base of 2CB-H and other potential things how you you purifying it?
walk me through the steps
id use ethanol personally, annhydrous hcl and etoh because the ONLY official solubility source based upon proven science on the internet i can find is 3mg/mL free base 2C-H in EtOH (no studies on the acetone, bunch forum talk here) however: https://pdf.benchchem.com/12644/Overcoming_solubility_issues_with_2C_B_Butterfly.pdf is good lab flowchart
Acetone and its miscibility with water ruins your day
Heres how i would see this: and id use ethanol like they do in that lab excerpt above. Using 2c-H as the example (Note: another user asked about this which is why i replied)


Scenario 1: Anhydrous Situation
Starting from 2C-H hydrochloride dissolved in anhydrous ethanol, and using a non-aqueous base (cant use NaOH (aq)) to keep the system anhydrous.

2C-H HCl solubility in Ethanol: 20 mg/mL (see https://www.caymanchem.com/product/11889/2c-h-hydrochloride)
2C-H Free Base: Miscible with alcohol (ethanol) and ether. Slightly soluble in water.

Anhydrous Basification & Salt Removal:
Preparation: In a dry flask equipped with a stir bar, dissolve 2C-H hydrochloride in anhydrous ethanol (using approximately 50 mL of ethanol per gram of 2C-H HCl, based on the 20 mg/mL solubility, or adjust for desired concentration). Ensure the system is kept anhydrous (e.g., with a drying tube).
Gradually add an anhydrous base like anhydrous K2CO3 powder to the stirred solution. This converts 2C-H HCl to its free base. Stir for sufficient time (e.g., 30-60 minutes) to ensure complete reaction.
Cool the mixture (e.g., in an ice bath) to minimize loss of the relatively volatile free base and to aid precipitation of the salt. Filter off the solid salt byproduct
Rinse the filter cake with a small amount of cold, anhydrous ethanol to recover any trapped 2C-H free base. Combine this rinse with the main filtrate. The filtrate now contains the 2C-H free base dissolved in anhydrous ethanol.
Concentration & Purification:
Remove the anhydrous ethanol using a form of evap. This will yield crude 2C-H free base. (Since 2C-H free base is miscible with alcohol, it will remain dissolved and be left behind as the ethanol evaporates).
If solid impurities are present, the crude free base might be dissolved in a minimal amount of a suitable solvent (e.g., diethyl ether or fresh anhydrous ethanol) and filtered again.


Scenario 2: Non-Anhydrous Situation (Water Present)
Starting from 2C-H hydrochloride dissolved in ethanol, and using aqueous NaOH (which introduces water).
Dissolve 2C-H hydrochloride in ethanol (using approximately 50 mL of ethanol per gram of 2C-H HCl).
Gradually add aqueous NaOH solution to the stirred ethanol solution until the pH is 10-12
This converts 2C-H HCl to its free base. Solid NaCl will form, and the system will now contain water, ethanol, NaCl, and 2C-H free base.
Filter off the solid NaCl
The filtrate contains 2C-H free base dissolved in a mixture of ethanol and water.
Transfer the ethanol/water/2C-H free base filtrate to a separatory funnel. Since 2C-H free base is "slightly soluble in water" but miscible with alcohol, it's beneficial to dilute the aqueous phase further with water if the ethanol concentration is too high, to encourage better partitioning into the organic extraction solvent. Add a water-immiscible organic solvent (e.g., diethyl ether or dichloromethane) to the separatory funnel.
Extract the 2C-H free base into the organic solvent by vigorous mixing and venting, followed by separation of layers. Repeat this extraction 2-3 times with fresh portions of the organic solvent. Combine all the organic extracts.
Dry the combined organic extracts using an anhydrous drying agent (Na2SO4 or MgSO4).
Filter off the spent drying agent.
Remove the organic solvent using a evap to obtain crude liquid 2C-H free base.
For high purity, consider vacuum distillation or recrystallization from a suitable solvent.
a. NaOH is BAD choice here due to its water content


If 2C Free Base has a melting point above the temperature you cool the solution to, then yes, it could THEORETICALLY crystallize.


your simply using acetone falls short


Is this all AI? This sounds like grok.

I don't even understand what you're talking about. Why are you talking about fucking with 2C-H?

Huh????

What's a drying tube.

Why are you doing this.
 
Is this all AI? This sounds like grok.

I don't even understand what you're talking about. Why are you talking about fucking with 2C-H?

Huh????

What's a drying tube.

Why are you doing this.
thank you! Yes its almost entirely AI and there are dozens of reasons it's obvious. Here's one. He says to cool the mixture to minimize loss of the relatively volatile freebase. 2C-H freebase boiling point is 160c at 10mm (.013% of standard atmospheric pressure) mercury vacuum. That is an extremely strong vacuum 2C-H is not going to just boil out of the flask. Even if this were a concern all of the heat created would first evaporate the ethanol...
AGAIN the title of this thread is How to Purify 2C-B at home. I personally live on Earth but maybe @jmc31337 lives on the moon where extremely high boiling freebase oils just evaporate freely or something.

This is what happens when you let AI answer all your questions. Grok as you call it.
At this point I think he trying to purposefully ruin the thread or appear like he knows something when it really destroys his entire credibility as a useful source of knowledge.

Fun fact NaOH does dissolve in many anhydrous alcohols including ethanol so it is acceptable as a base while K2CO3 is completely insoluble...

"remove the organic solvent using a evap to obtain crude liquid 2C-H free base. For high purity, consider vacuum distillation or recrystallization from a suitable solvent" - Good luck recrystallizing 2C-H freebase which is an oil.

It's easy to recognize when someone tries to use AI to explain a lot of things because it just comes out as totally impractical garbage when compared to real world situations.
I'm sure a lot of us on BL would appreciate if people stopped using AI to explain things that they have zero first hand experience. If I come off as an asshole for saying that then so be it. I personally find it frustrating and just plain sad.

A drying tube is a piece of lab glass usually filled with calcium chloride and cotton to prevent atmospheric moisture from entering the system. It's almost like he thinks enough atmospheric moisture
 
Last edited:
thank you! Yes its almost entirely AI and there are dozens of reasons it's obvious. Here's one. He says to cool the mixture to minimize loss of the relatively volatile freebase. 2C-H freebase boiling point is 160c at 10mm (.013% of standard atmospheric pressure) mercury vacuum. That is an extremely strong vacuum 2C-H is not going to just boil out of the flask. Even if this were a concern all of the heat created would first evaporate the ethanol...
AGAIN the title of this thread is How to Purify 2C-B at home. I personally live on Earth but maybe @jmc31337 lives on the moon where extremely high boiling freebase oils just evaporate freely or something.

This is what happens when you let AI answer all your questions. Grok as you call it.
At this point I think he trying to purposefully ruin the thread or appear like he knows something when it really destroys his entire credibility as a useful source of knowledge.

Fun fact NaOH does dissolve in many anhydrous alcohols including ethanol so it is acceptable as a base while K2CO3 is completely insoluble...

"remove the organic solvent using a evap to obtain crude liquid 2C-H free base. For high purity, consider vacuum distillation or recrystallization from a suitable solvent" - Good luck recrystallizing 2C-H freebase which is an oil.

It's easy to recognize when someone tries to use AI to explain a lot of things because it just comes out as totally impractical garbage when compared to real world situations.
I'm sure a lot of us on BL would appreciate if people stopped using AI to explain things that they have zero first hand experience.

A drying tube is a piece of lab glass usually filled with calcium chloride and cotton to prevent atmospheric moisture from entering the system. It's almost like he thinks enough atmospheric moisture
nope no Ai as Ai especially grok wont tell ya how purify 2C compounds

lets recap since i did the homework for you

1. I came in here not knowing how to synthesize 2c-b 2c-h etc, which is fine since this was about purification
2. I stated acetone is the organic chem basic WASH not purfication - that started all this back n forth
3. Then someone, says in side path: how we get annhydrous HCl and i answered that with backyard clandestine methods chloro-alki - we need HCl annhydrous clearly from the above
4. Then someone says some crap about acetone being used to further form crystals and precipitations (could be as an ANTISOLVENT)
5. The i realize for starters the thread mentioned prufiy but never mentioned what the suspect contaminant was so I threw some basic kitchen crap as example (sugar, aspartame caffiene)
6. We then discussed how NONE of us began the conversation with which form freebase or salt we were even dealing with
7. Then we tossed about some research papers, and looked for solvents with KNOWN solubility to 2c-b 2c-h
8. 2c-h became another side path question
9. I then jsut gave you the answer and mentioned EtOH annhydrous (bcuz again lil moonshine backyard diy goes long way) opposed to the other cited solubility to the other chemicals
10. I then showed ya the proper way to purify by taking the freebase and the salt back to start and reformation of the precip
11. Then we went on a tangent of crystallography and deep dove on that, none of us can determine the temp at any of these compounds that crystallization begins
12. So i included the flow chart (to avoid synthesis rule braking,) in case one yall that thinks they know something about acetone could measure that for us and be done with it
13. I also said that i wouldnt use NaOH and get that cakey pasty stuff but used it as example, fun fact plenty of other anhhydrous bases exist but im doing your homework, in fact i would go full annhydrous diy backyard style
14. Lastly, cool the boiling point and nowhere do my final responses to ya dog ate your homework replies all through out do i mention anything about heating this up? Nowhere
And, what i would use as guidance from the lab texts to form crystalls is melting points not boiling points since this compound being liquid at STP

gave ya the answer except the final annhydrous gasssing and the final mL EtOH for the 2C-B which is easily calculated based upon KNOWN solubility, because i ddint wanna get kicked outta the forum (but its around 10+mL id start since its 3mg/mL)
no heat mentioned just evap (vacuum can do that without heat,) and actually my method uses ice baths

---------
Btw if we just go done saying cooled temps for solubility to crystallize why would we do boiling points? but ya made a fair EVAP the EtOH point


Last minute edit: storage and handling, we know from shulgins' reports this stuff oxidizes in the atmosphere around us, so while we argue about this freebase we're losing product to time
Methanol, now we need get some that bcuz itll dissolve both hcl and freebase nicely and boils at 148C meaning i think based upon YOUR boiling point chart, may as well use it right?
we can do the ole vet cat tranq hot plate method, place lil bit solution on pyrex glass heat it lil and if the hcl was dissolved we get the hcl back
if its freebase, we're back to the annhydrous

If ya handed it oft to someone, we're now back to them hitting this forum asking i just got 2c-b / 2c-h how do i purify it

round n round we go
 
Last edited:
11. Then we went on a tangent of crystallography and deep dove on that, none of us can determine the temp at any of these compounds that crystallization begins
Actually I can it's called using a data logging thermometer. In fact while you're recording the temperature during the cooldown, you'll actually see an exothermic event the moment crystals start to form in solution. You KNOW it's done when the temperature change over time resumes the same slope as before the exothermic moment.

When I heard him say Grok i literally thought he meant garbage as some sort of european slang.
I think you should probably stick to doing your own homework first cuz you must have failed high school freshmen chemistry.

You tell me since you're the one (not ai) who mentioned 2C-H being relatively volatile...

In chemistry, volatility describes how easily a substance turns into a gas or vapor. [1]

Key Characteristics
  • Low Boiling Point: Volatile substances require very little heat to break their intermolecular bonds and evaporate.
  • High Vapor Pressure: They have a high vapor pressure, meaning their molecules easily escape from the surface of a liquid or solid into the air.
 
Last edited:
Actually I can it's called using a data logging thermometer. In fact while you're recording the temperature during the cooldown, you'll actually see an exothermic event the moment crystals start to form in solution. You KNOW it's done when the temperature change over time resumes the same slope as before the exothermic moment.

When I heard him say Grok i literally thought he meant garbage as some sort of european slang.
I think you should probably stick to doing your own homework first cuz you must have failed high school freshmen chemistry.

You tell me since you're the one (not ai) who mentioned 2C-H being relatively volatile...

In chemistry, volatility describes how easily a substance turns into a gas or vapor. [1]

Key Characteristics
  • Low Boiling Point: Volatile substances require very little heat to break their intermolecular bonds and evaporate.
  • High Vapor Pressure: They have a high vapor pressure, meaning their molecules easily escape from the surface of a liquid or solid into the air.
put up here then, run the lab experiment and put a digit up there
like i said the only ral data comes from that solubility from the lab link above
im done here the answer sits at EtOH and thats sound bcuz thats what the flow chart lab used as well

and it so happens to have a solubility number tried and true (whereas everything else was all speculation from yall)


done here movin on
 
My friend has some 2C-B wants to purify it at home. Despite the username he's not Walter White.

Is it possible to do with water or alcohol?

It's supposedly HCL poweder. Any tips?
purify? why does he assume it isn't already pure?

look up acid-base washes, recrystallization & acetone purification teks to understand how;
 
thank you! Yes its almost entirely AI and there are dozens of reasons it's obvious. Here's one. He says to cool the mixture to minimize loss of the relatively volatile freebase. 2C-H freebase boiling point is 160c at 10mm (.013% of standard atmospheric pressure) mercury vacuum. That is an extremely strong vacuum 2C-H is not going to just boil out of the flask. Even if this were a concern all of the heat created would first evaporate the ethanol...
AGAIN the title of this thread is How to Purify 2C-B at home. I personally live on Earth but maybe @jmc31337 lives on the moon where extremely high boiling freebase oils just evaporate freely or something.

This is what happens when you let AI answer all your questions. Grok as you call it.
At this point I think he trying to purposefully ruin the thread or appear like he knows something when it really destroys his entire credibility as a useful source of knowledge.

Fun fact NaOH does dissolve in many anhydrous alcohols including ethanol so it is acceptable as a base while K2CO3 is completely insoluble...

"remove the organic solvent using a evap to obtain crude liquid 2C-H free base. For high purity, consider vacuum distillation or recrystallization from a suitable solvent" - Good luck recrystallizing 2C-H freebase which is an oil.

It's easy to recognize when someone tries to use AI to explain a lot of things because it just comes out as totally impractical garbage when compared to real world situations.
I'm sure a lot of us on BL would appreciate if people stopped using AI to explain things that they have zero first hand experience. If I come off as an asshole for saying that then so be it. I personally find it frustrating and just plain sad.

A drying tube is a piece of lab glass usually filled with calcium chloride and cotton to prevent atmospheric moisture from entering the system. It's almost like he thinks enough atmospheric moisture

Grok is a LLM bot that is friendly to drug work. It also makes CSAM.

"I personally live on Earth but maybe @jmc31337 lives on the moon where extremely high boiling freebase oils just evaporate freely or something."

Yes, they are on a datacenter in space. BTW the sentient space laser episode from Cowboy Bebop was great.

Anyways. Seriously, what a fucking clown. Why even bother. Bad information. Another bloke blocked.
 
Top